> For the complete documentation index, see [llms.txt](https://help.connected.illumina.com/llms.txt). Markdown versions of documentation pages are available by appending `.md` to page URLs; this page is available as [Markdown](https://help.connected.illumina.com/emedgene/emedgene-analyze-manual/variant_page/summary_section.md).

# Summary tab

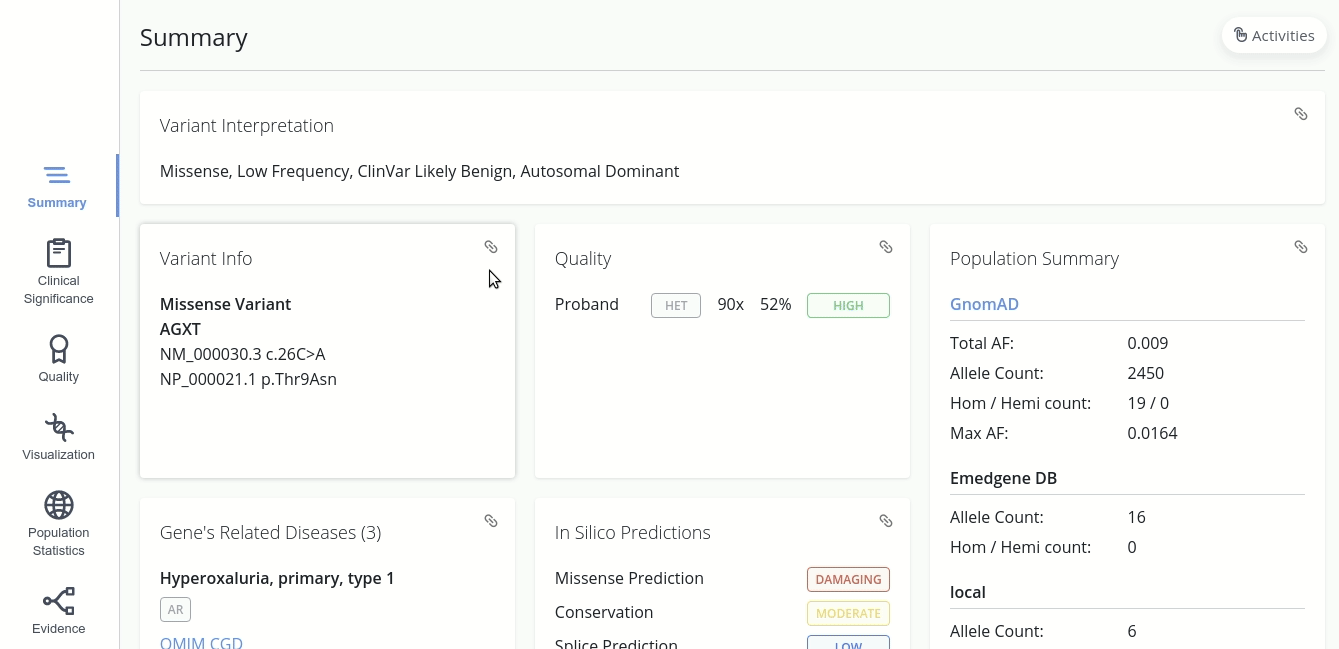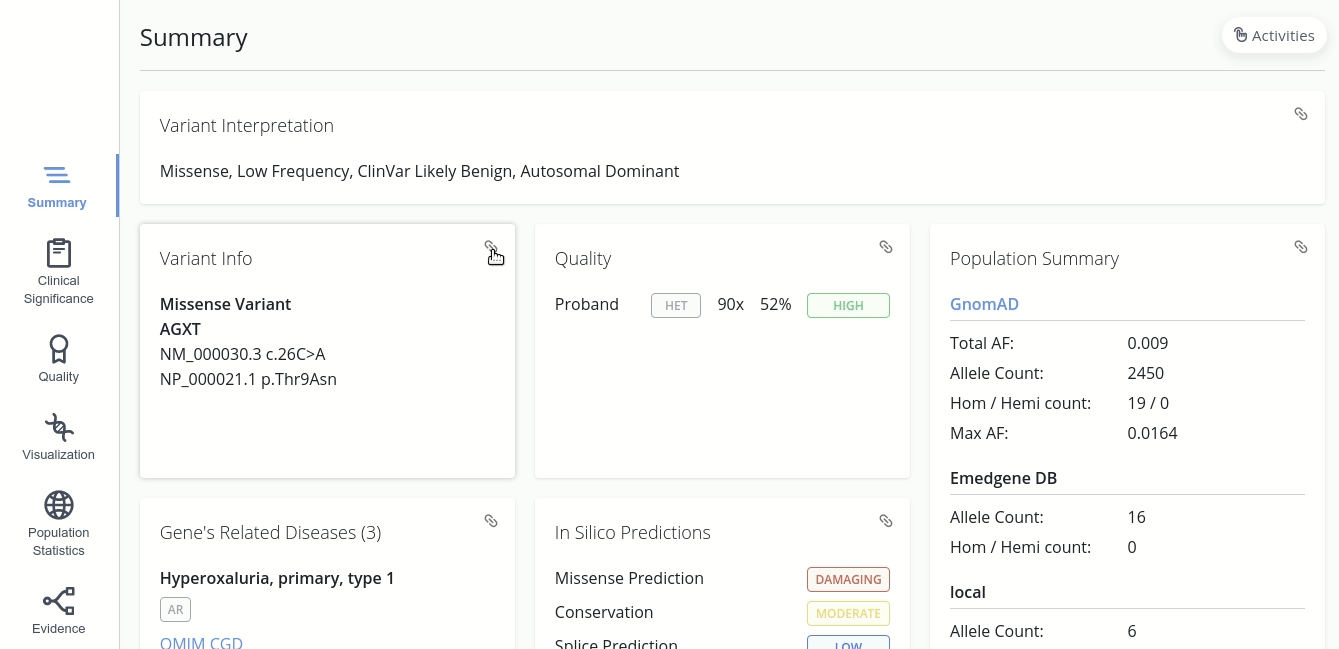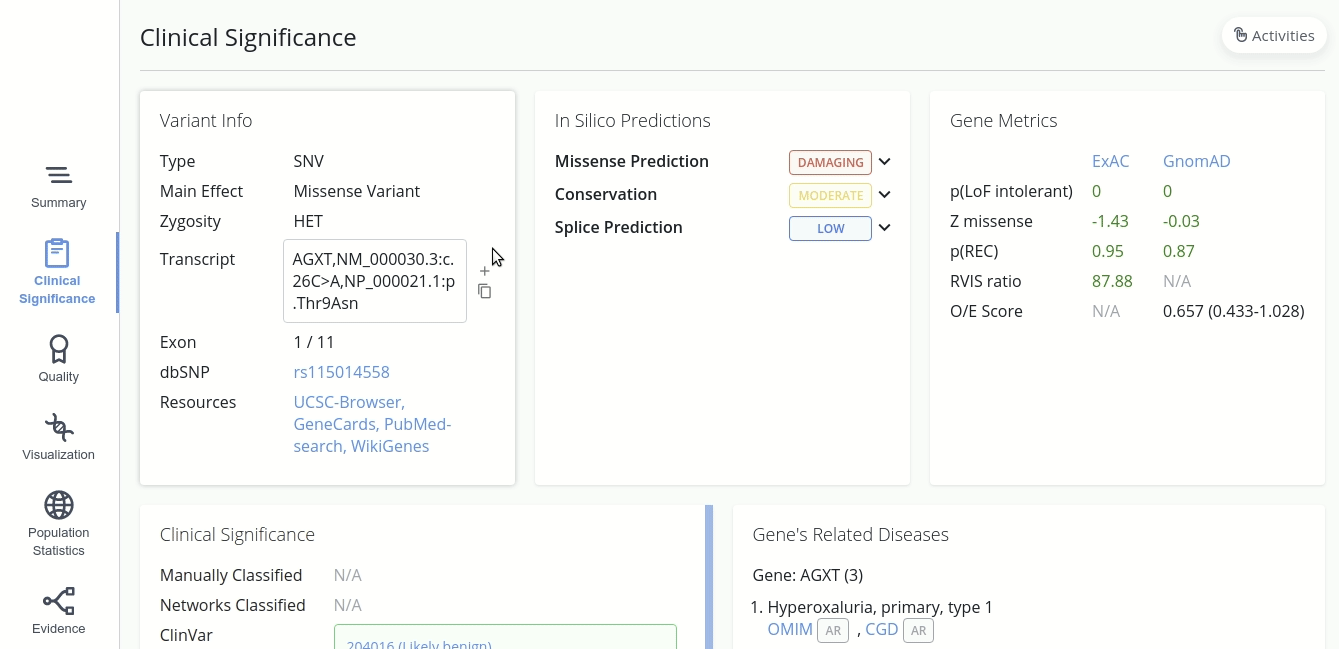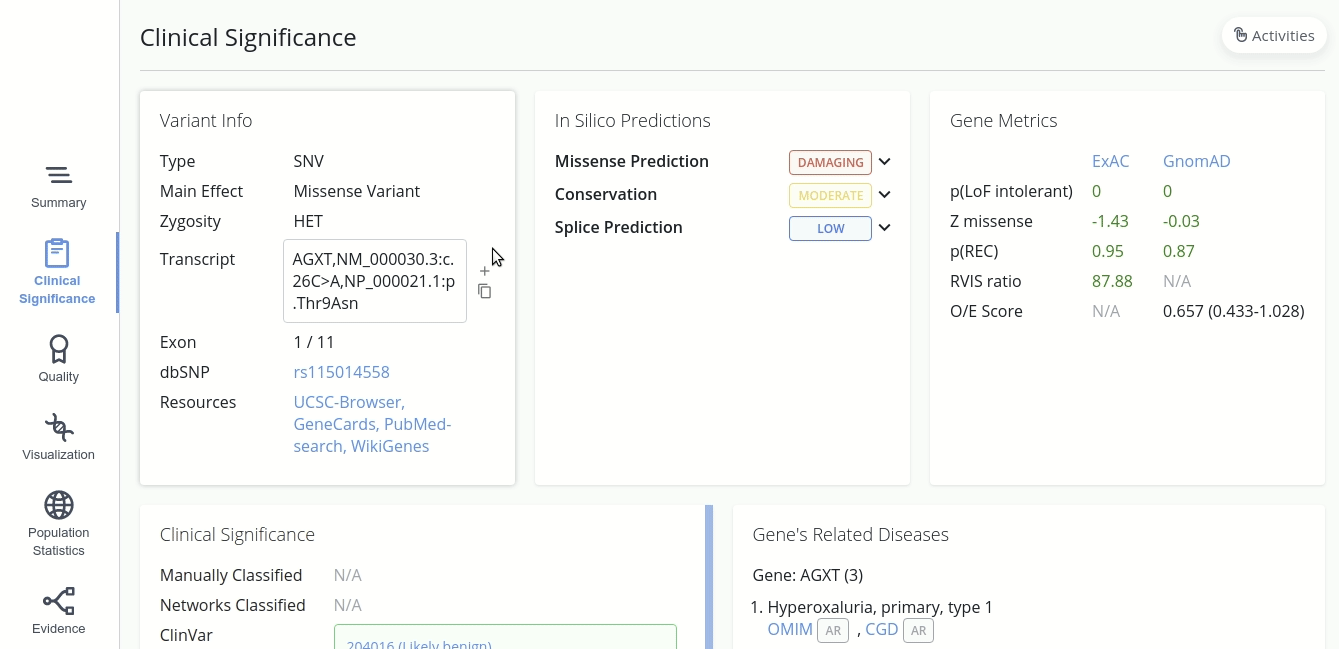
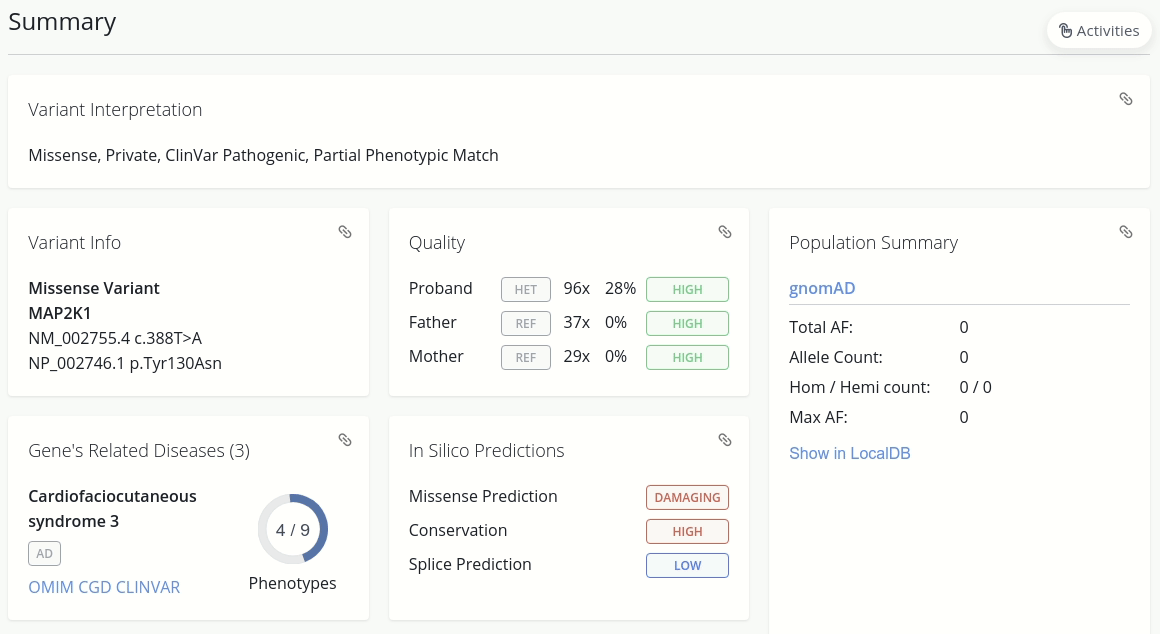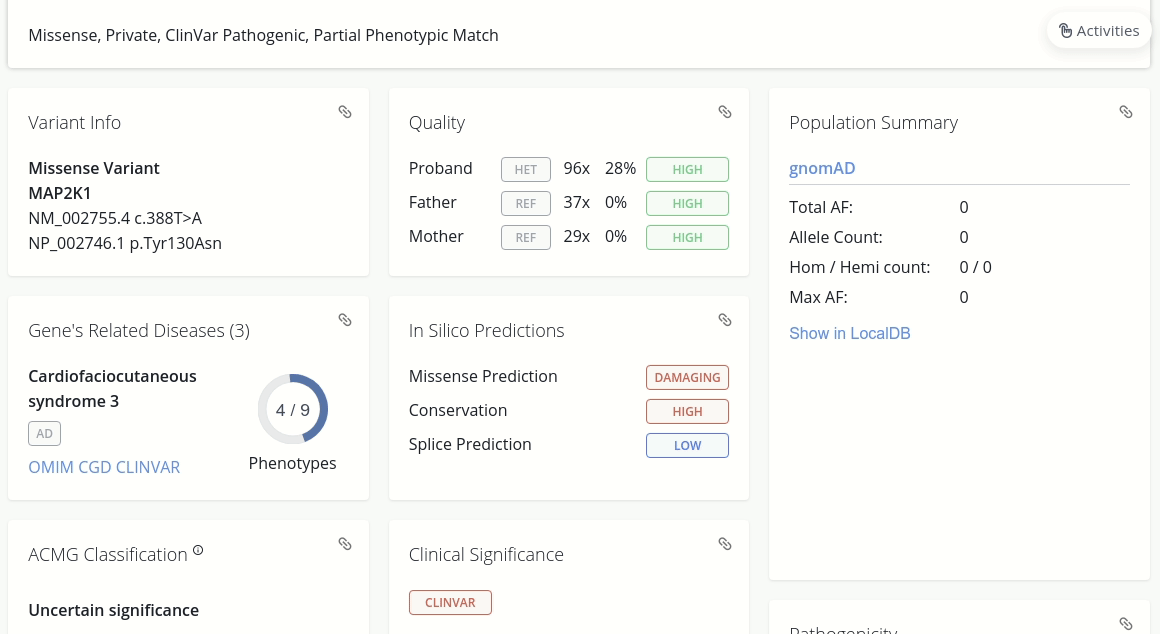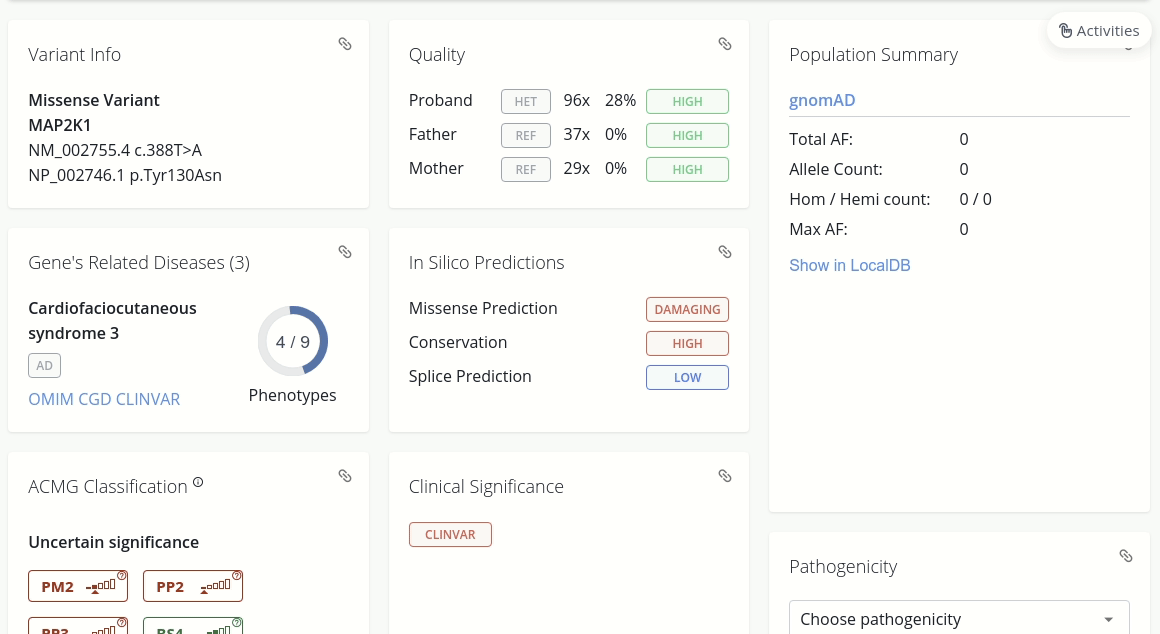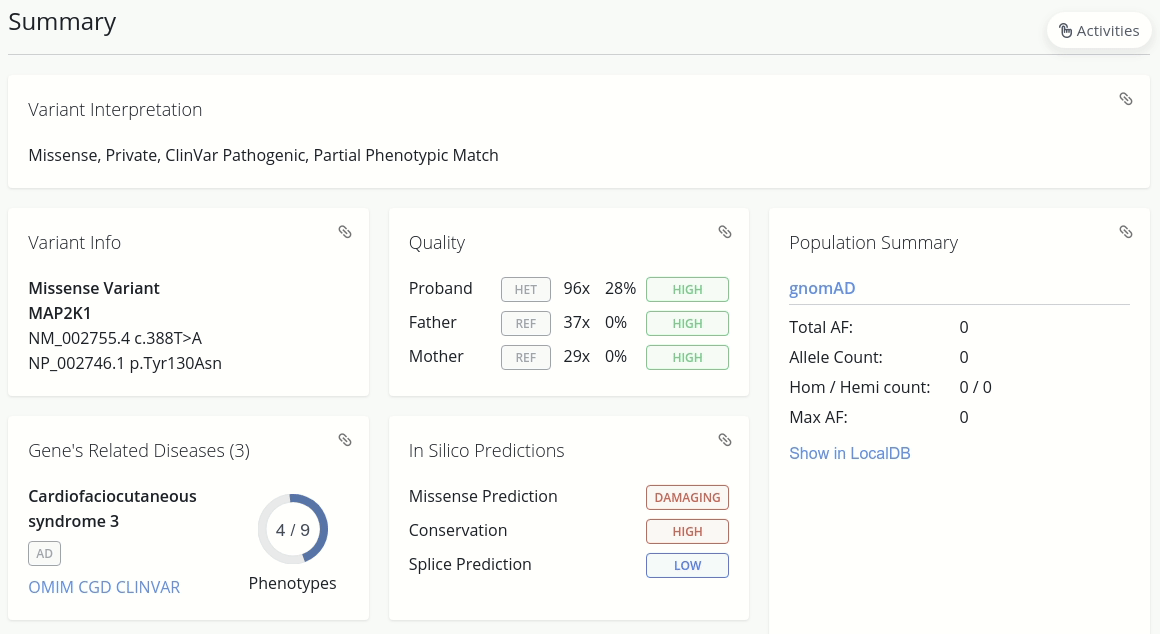
The **Summary** tab highlights core variant-related information from other [**Variant page**](/emedgene/emedgene-analyze-manual/variant_page/variant_page.md) tabs:

* [**Clinical Significance tab**](/emedgene/emedgene-analyze-manual/variant_page/clinical_significance_section.md)**:** [**Variant Info**](#variant-info), [**In silico Prediction**](#in-silico-prediction), [**Gene's related diseases**](#genes-related-diseases), [**Clinical significance**](#clinical-significance) **cards**
* [**Quality tab**](/emedgene/emedgene-analyze-manual/variant_page/quality_section.md)**:** [**Quality card**](#quality)
* [**Connected variants** **tab**](/emedgene/emedgene-analyze-manual/variant_page/connected-variants-tab.md)**: Connected variants card**
* [**Population Statistics** **tab**](/emedgene/emedgene-analyze-manual/variant_page/population_statistics_section.md)**:** [**Population Summary card**](#population-summary)
* [**Evidence tab**](/emedgene/emedgene-analyze-manual/variant_page/evidence_section.md): [**Variant Interpretation**](#variant-interpretation), [**ACMG Classification**](#acmg-classification), [**Pathogenicity**](#pathogenicity) **cards**

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-28c1243deba01fc75fca84a8d01a0fe8ff9ac48e%2Fsummary_section_1_summary%2Bwithout%2Bhgmd.gif?alt=media" alt=""><figcaption><p><strong>Summary tab</strong> overview.</p></figcaption></figure></div>

Each **Summary tab** card links to its original location on the **Variant page** where you can review full details.

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-a91c9c0a015b0f8b475e764f48e4be545fe1834d%2Fsummary_section_2.gif?alt=media" alt=""><figcaption><p><strong>Summary tab cards</strong> link to their source tabs.</p></figcaption></figure></div>

Let’s look more closely at each card.

***

## Variant Interpretation

Notes are added automatically or manually in the [**Evidence** **tab**](/emedgene/emedgene-analyze-manual/variant_page/evidence_section.md). They appear only when present.

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-e7a4ec534506986b04213a4ba452f2b69d7f9286%2Fsummary_section_3.png?alt=media" alt=""><figcaption><p><strong>Variant Interpretation card</strong>.</p></figcaption></figure></div>

***

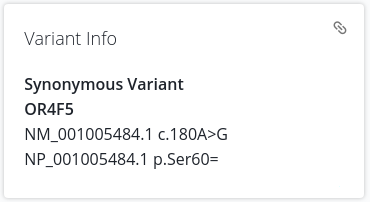
## Variant Info

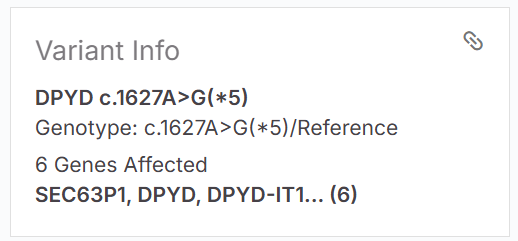
**SNV/Indel, MNV (v100.39+), mtDNA SNV/Indel**

{% columns %}
{% column width="50%" valign="middle" %}

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-983e402a4c28e4e0067b0405e0b53732571ecc90%2Fsummary_section_4.png?alt=media" alt=""><figcaption></figcaption></figure></div>
{% endcolumn %}

{% column width="50%" %}

* Main effect
* Gene symbol
* HGVS descriptions at the coding DNA and protein levels
  {% endcolumn %}
  {% endcolumns %}

**DEL, DUP, LOH/ROH**

{% columns %}
{% column width="50%" valign="middle" %}

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-07b041dd80950d8ebdc1861cdf84eee812efa2e8%2Fsummary_section_5.png?alt=media" alt=""><figcaption></figcaption></figure></div>
{% endcolumn %}

{% column width="50%" %}

* Variant length
* Variant type
* Number of genes involved
* Gene symbol(s)

{% hint style="info" %}
Hover to see the full gene list.
{% endhint %}
{% endcolumn %}
{% endcolumns %}

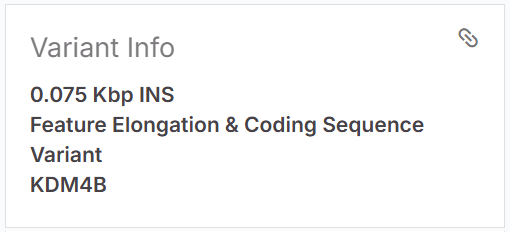
**INS**

{% columns %}
{% column width="50%" valign="middle" %}

<figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-13444c84388a243b1f337b152c92a852efb963f9%2Fimage%20(186).png?alt=media" alt=""><figcaption></figcaption></figure>
{% endcolumn %}

{% column width="50%" %}

* Variant length
* Variant type
* Main effect
* Gene symbol
  {% endcolumn %}
  {% endcolumns %}

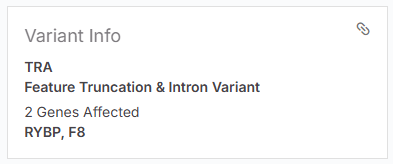
**TRA (v100.41+)**

{% columns %}
{% column width="50%" valign="middle" %}

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-681c3c989167df38487ade5886cdd5906948fea3%2Ftra.png?alt=media" alt="Variant Info card for a translocation"><figcaption></figcaption></figure></div>
{% endcolumn %}

{% column width="50%" %}

* Variant type
* Main effect
* Number of genes involved
* Gene symbols
  {% endcolumn %}
  {% endcolumns %}

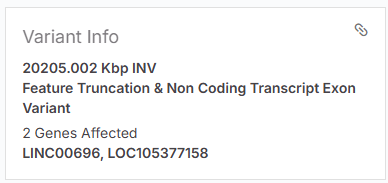
**INV (v100.41+)**

{% columns %}
{% column width="50%" valign="middle" %}

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-ebd964c62e91bdfe56d3719e3e852cd589be0c36%2Finv.png?alt=media" alt="Variant Info card for an inversion"><figcaption></figcaption></figure></div>
{% endcolumn %}

{% column width="50%" %}

* Variant length
* Variant type
* Main effect
* Number of genes involved
* Gene symbol(s)
  {% endcolumn %}
  {% endcolumns %}

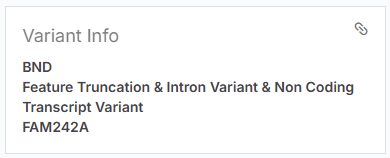
**BND (v100.41+)**

{% columns %}
{% column width="50%" valign="middle" %}

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-3e6b50781470f8ead523d73428479d0357ed3f52%2Fbnd1.png?alt=media" alt="Variant Info card for a breakend"><figcaption></figcaption></figure></div>
{% endcolumn %}

{% column width="50%" %}

* Variant type
* Main effect
* Gene symbol
  {% endcolumn %}
  {% endcolumns %}

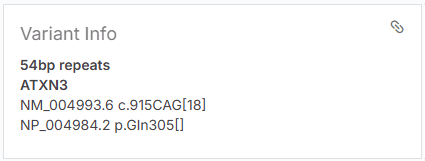
**STR**

{% columns %}
{% column width="50%" valign="middle" %}

<figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-8cad0e613ed40f8f081b5b628d05f00edf8410f4%2Fimage%20(190).png?alt=media" alt=""><figcaption></figcaption></figure>
{% endcolumn %}

{% column width="50%" %}

* Total length of the repeat region
* Gene symbol
* HGVS descriptions at the coding DNA and protein levels
  {% endcolumn %}
  {% endcolumns %}

**Haplotype**

{% columns %}
{% column width="50%" valign="middle" %}

<figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-c5afccdf474b9cb77f20788c3b9a978d9b76ba81%2Fimage%20(192).png?alt=media" alt=""><figcaption></figcaption></figure>
{% endcolumn %}

{% column width="50%" %}

* Haplotype name
* Individual genotype
* Number of genes involved
* Gene symbol(s)
  {% endcolumn %}
  {% endcolumns %}

***

## Quality

**SNV/Indel, MNV (v100.39+), mtDNA SNV/Indel, STR**

{% columns %}
{% column width="50%" valign="middle" %}

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-06432631921f2c5a2ea9ff519ad3f55ddd72290f%2Fsummary_section_6_summary%2Bsection%2Bsnv.png?alt=media" alt="" width="405"><figcaption></figcaption></figure></div>
{% endcolumn %}

{% column width="50%" %}

* Variant caller badge
* Zygosity per sample
* Depth of coverage per sample
* Percentage of alternative allele reads per sample
* Overall variant quality per sample
  {% endcolumn %}
  {% endcolumns %}

**DEL, DUP, LOH/ROH, INS, TRA (v100.41+), INV (v100.41+), BND (v100.41+), Haplotype**

{% columns %}
{% column width="50%" valign="middle" %}

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-fe2c5ce56e950fcbd6795901c95f6531ac31788f%2Fsummary_section_7_summary%2Bsection%2Bcnv.png?alt=media" alt="" width="408"><figcaption></figcaption></figure></div>
{% endcolumn %}

{% column width="50%" %}

* Variant caller badge
* Zygosity per sample
* Overall variant quality per sample
  {% endcolumn %}
  {% endcolumns %}

***

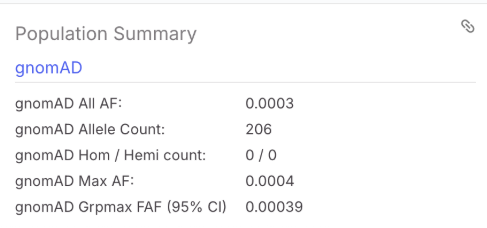
## Population Summary

**Population Summary card** features:

* Link to the gnomAD variant entry for the specific dataset used for annotation.
* Population statistics from a combined exome + genome gnomAD dataset:
  * **gnomAD All AF**: Overall alternative allele frequency across gnomAD populations.
  * **gnomAD Allele count**: Number of observed alternate alleles in the gnomAD dataset.
    * For mtDNA variants, **Homoplasmy Count** is shown instead.
  * **gnomAD Hom/Hemi count**: Number of gnomAD subjects who are homozygous or hemizygous for this variant.
    * For mtDNA variants, **Heteroplasmy Count** is shown instead.
  * **gnomAD Max AF**: The highest alternative allele frequency of the variant among gnomAD ancestry groups. For each ancestry group, Emedgene first combines gnomAD exome and genome counts, then takes the highest combined frequency: `(exome allele count + genome allele count) / (exome allele number + genome allele number)`. There is no minimum sample-size (AN) cutoff: a group is used even if it was genotyped in 1000 alleles or fewer.\
    **Note:** Not to be confused with **Max AF (%)** in the **Variant table**, which is the highest frequency among all public population databases. In that column, gnomAD exomes and gnomAD genomes are compared separately, and an ancestry group is included only if AN > 1000.
  * **gnomAD Grpmax FAF (95% CI)** (v100.41+): The highest [filtering allele frequency](https://gnomad.broadinstitute.org/help/faf) observed across ancestry groups in a combined exome/genome gnomAD dataset. It uses a 95% confidence interval.
    * Available for SNV, indel, and MNV variants annotated with the [gnomAD 4.1 All](/emedgene/emedgene-analyze-manual/settings/organization_settings_-330+/workbench-and-pipeline/gnomad-annotation-v100.41+.md) dataset.
* Population statistics from organization databases, if available:
  * **Allele count**: Count of alternative alleles.
  * **Hom/Hemi count**: Count of alternative alleles in a homozygous or hemizygous state.

The card is available for SNV/Indel, MNV (v100.39+), mtDNA SNV/Indel, DEL, DUP, INS, TRA (v100.41+), INV (v100.41+) variant types.

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-c2493796ed82c4b96e1e8db028f25565dd9457c7%2Fpop%20sum.png?alt=media" alt=""><figcaption><p><strong>Population Summary card</strong>.</p></figcaption></figure></div>

***

### STR repeats distribution

For STR variants, **STR Repeats Distribution** replaces the **Population Summary card**. It displays allele counts from gnomAD and 1000 Genomes, and gnomAD pathogenicity ranges.

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-b421a405ffa838439eb2288418e8e885f8a15ee4%2Fsummary_section_9_summary_section_str_repeats_dist.png?alt=media" alt=""><figcaption><p><strong>STR repeats distribution card</strong>.</p></figcaption></figure></div>

***

## Gene's related diseases

The card features:

* Number of gene–disease connections in the Emedgene knowledge base
* Name of the selected disease
* Inheritance mode(s) of the selected disease
* GenCC validity badge (100.39+)\
  When the selected disease has one or more GenCC (Gene Curation Coalition) database entries, a badge appears next to its inheritance mode. The badge represents the highest gene–disease validity classification.\
  If multiple GenCC entries exist for the same gene–disease connection, a “+n” indicator appears beside the badge. Hover over the indicator to view additional classifications and their sources.
* Links to the gene–disease connection source(s) for the selected disease:
  * OMIM
  * Academic papers in the Emedgene knowledge graph
  * CGD
  * ClinVar
  * Orphanet
  * GenCC (100.39+)

{% hint style="info" %}
**Note:**

GenCC is included as a gene–disease source starting with [Emedgene knowledge base version 82](/emedgene/release-notes/knowledgebase_updates/2025/zoidberg-82-21st-october-2025.md).

To view GenCC annotations in the UI, you must use [platform version 100.39](/emedgene/release-notes/workbench-and-pipeline-updates/new-in-emedgene-v100.39.0-october-16th-2025.md) or later.

On older platform versions, the GenCC validity badge and submitter details are not displayed, and GenCC links are inactive.
{% endhint %}

* Link to the disease page in MONARCH (100.39+)
* Phenotype match summary (displayed only for tagged variants):
  * Number of disease phenotypes that match the patient’s phenotypes, out of the total
  * List of disease phenotypes with indicated [match levels](/emedgene/emedgene-analyze-manual/reviewing_a_case/phenotypic-match/phenotypic_match_strength.md)

{% hint style="warning" %}
**TRA (v100.41+), INV (v100.41+), BND (v100.41+):** Gene-disease annotation for these variant types considers genes at the breakpoint position(s) where applicable.
{% endhint %}

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-179f13c8a9598f03d53ff601747e5427cb9f5500%2Fimage%20(95).png?alt=media" alt=""><figcaption><p><strong>Gene's related diseases card.</strong></p></figcaption></figure></div>

***

## Pathogenicity

View the user-assigned **Pathogenicity**. Change it or select a value from the dropdown when it is empty.

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-80316148e22badc714efc553f2a4577143d29d59%2Fsummary_section_11.png?alt=media" alt="" width="269"><figcaption><p><strong>Pathogenicity card.</strong></p></figcaption></figure></div>

***

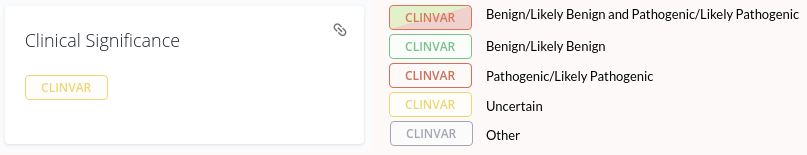
## Clinical significance

This card highlights existing pathogenicity classifications in public and private variant databases, including [**Curate**](/emedgene/emedgene-curate-manual/curate_variants.md). Each source has one badge. **Uncertain** and **Other** classifications appear only when that database has no **Benign/Likely Benign** or **Pathogenic/Likely Pathogenic** classifications for the variant.

The card is available for SNV/Indel, MNV (v100.39+), mtDNA SNV/Indel, DEL, DUP, INS, TRA (v100.41+), INV (v100.41+), STR, haplotype variant types.

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-395046886ec0e15aac4c28982bfdd144b754c2ea%2Fsummary_section_12_image.png?alt=media" alt=""><figcaption><p><strong>Clinical significance card.</strong></p></figcaption></figure></div>

***

## ACMG Classification

Shows ACMG tags assigned to the variant and the resulting ACMG pathogenicity class and score.

The card is available for SNV/Indel, mtDNA SNV/Indel, DEL, DUP variant types.

**SNV/Indel, mtDNA SNV/Indel**

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-650729d01526d93e37fec188e2c59f1eec2e2587%2Fsummary_section_14_Screenshot_20.png?alt=media" alt="" width="406"><figcaption><p><strong>ACMG Classification card</strong> for sequence and mtDNA variants.</p></figcaption></figure></div>

**DEL, DUP**

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-f990c5ddc7a3fa5513b5066692867dbcea76f0e3%2Fsummary_section_15.png?alt=media" alt="" width="410"><figcaption><p><strong>ACMG Classification card</strong> for CNVs.</p></figcaption></figure></div>

***

## In Silico Prediction

Overall estimates of *in silico* prediction results: **Missense Prediction**, **Conservation**, and **Splicing Prediction**.

Availability of a specific prediction category depends on variant type:

* **Small variant (SNV/Indel/MNV):** **Missense Prediction**, **Conservation**, and **Splicing Prediction**
* **mtDNA (SNV/Indel):** **Missense Prediction, Conservation**
* **Other variant types**: Not available

<div align="left"><figure><img src="https://1131024994-files.gitbook.io/~/files/v0/b/gitbook-x-prod.appspot.com/o/spaces%2FGCW0DnLlE7QjoZPNmKIi%2Fuploads%2Fgit-blob-a19e732d87a83405799a4b79c5287bd8ce5e1b09%2Fsummary_section_16.png?alt=media" alt=""><figcaption><p><strong>In Silico Prediction card</strong>.</p></figcaption></figure></div>


---

# Agent Instructions
This documentation is published with GitBook. GitBook is the documentation platform designed so that both humans and AI agents can read, navigate, and reason over technical content effectively. Learn more at gitbook.com.

## Querying This Documentation
If you need additional information that is not directly available in this page, you can query the documentation dynamically by asking a question.

Perform an HTTP GET request on the current page URL with the `ask` query parameter, and the optional `goal` query parameter:

```
GET https://help.connected.illumina.com/emedgene/emedgene-analyze-manual/variant_page/summary_section.md?ask=<question>&goal=<endgoal>
```

`ask` is the immediate question: it should be specific, self-contained, and written in natural language.
`goal` is optional and describes the broader end goal you are ultimately trying to accomplish on behalf of the user. GitBook uses it to tailor the answer towards what is most useful for that goal.

The response will contain a direct answer to the question and relevant excerpts and sources from the documentation.

Use this mechanism when the answer is not explicitly present in the current page, you need clarification or additional context, or you want to retrieve related documentation sections.